Sterilization Packaging: A Focused Framework for Sterilant Access and Package Integrity
Scope: two packaging questions
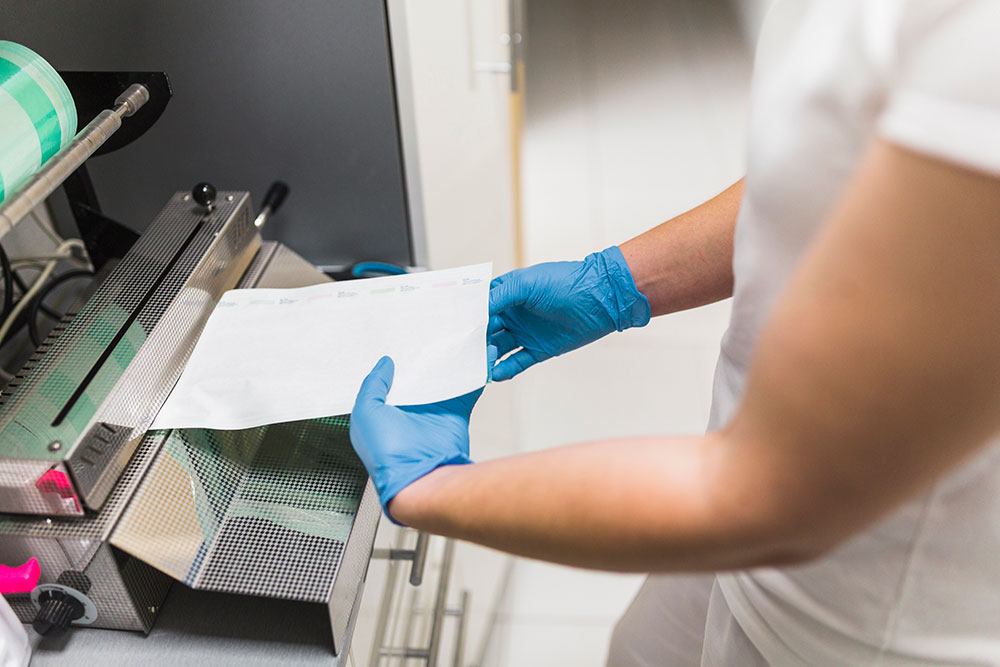
Sterilization packaging has two related but distinct roles: it must support the selected sterilization process, and it must preserve the sterile barrier until the package is opened for use. A package that protects well but restricts sterilant contact may interfere with processing. A package that permits exposure but is damaged, poorly sealed, or mishandled may not protect the processed contents afterward. Packaging should therefore be evaluated as one part of the complete reprocessing system rather than as an isolated wrapping task. [1][2]
This article focuses on the relationship between sterilant access, package configuration, and package integrity. It is intended to complement, not replace, separate guidance on package–sterilizer compatibility and event-related sterile-barrier inspection. Those topics may overlap in practice, but this framework uses them to organize a focused review: first, whether the process can reach the device; second, whether the finished package remains an effective barrier afterward.
The discussion does not establish detailed requirements for wet-pack management, cooling or transfer practices, heat-seal monitoring, shelf life, or change-control procedures. It also does not establish requirements for every reusable CSSD packaging scenario. Device, packaging, sterilizer, and facility instructions remain controlling.
Question 1: Can the sterilant reach the device?
Packaging materials and configurations should be compatible with the selected sterilization method and cycle. The package should permit the relevant sterilant to reach the surfaces requiring treatment while supporting removal of air, moisture, or residual sterilant as appropriate for the process. Device geometry, folds, lumens, package density, and the arrangement of multiple packages can affect access. [1][2]
CDC recommendations state that packaging materials should be compatible with the sterilization process and, where applicable, cleared by the U.S. Food and Drug Administration. Selection and preparation should also follow the device manufacturer’s reprocessing instructions and the packaging manufacturer’s instructions for use. [3]
Compatibility is not determined by material appearance alone. A transparent pouch, nonwoven wrap, rigid container, or specialty tray may have different limitations related to penetration, drying, sealing, weight, or configuration. A packaging system may be intended for a sterilization method yet perform differently when used outside its validated or instructed configuration.
Changes in pouch size, wrap technique, tray density, device orientation, or load arrangement can alter the challenge presented to the process. CDC recommendations identify packaging, loading, and packaging-material selection as factors to consider when sterilization monitoring indicates a problem. [3]
Question 2: Can the package maintain the sterile barrier?
After processing, the package must withstand the handling, transport, storage, and environmental stresses expected before point of use. Punctures, tears, delamination, seal defects, abrasion, crushed corners, and contamination introduced during handling can compromise the sterile barrier. Inspection of damaged packages is therefore a separate question from whether a chemical indicator shows exposure to process conditions. [1][2]
Inspection should consider the package as a finished system, including its material, seals, closures, and assembly. The FDA’s recognized-standards listing identifies ISO/TS 16775 as informative guidance for applying ISO 11607-1 and ISO 11607-2. ISO 11607-1 and ISO 11607-2 concern packaging systems for terminally sterilized medical devices: the first addresses materials and sterile-barrier systems, and the second addresses validation of forming, sealing, and assembly processes. These standards and the FDA listing should not be presented as direct, universal requirements for every reusable CSSD packaging situation. [4]
The distinction remains useful for quality review. Package integrity depends both on the selected components and on how the finished package is formed, sealed, assembled, handled, and stored. The applicable instructions and facility procedures determine how those principles are applied in a particular CSSD.
A focused CSSD review framework
Begin with the device and process
Confirm the device manufacturer’s reprocessing instructions and the packaging manufacturer’s instructions for use before selecting or preparing a package. Review features that may affect packaging and sterilant access, such as sharp edges, hinges, concave surfaces, lumens, removable components, and moisture-retaining areas. Confirm that the packaging system is intended for the sterilization method and cycle used by the facility.
Do not assume that a package is suitable solely because it is commonly used or visually transparent. The applicable instructions should determine the permitted configuration, preparation method, loading approach, and relevant limitations. Packaging-system information may support this review, but it does not replace device-specific or facility-specific instructions.
Evaluate configuration, not only material
When reviewing a package or investigating a recurring issue, ask:
- Was the device prepared and arranged according to its instructions for use?
- Were hinges opened and removable components separated when required?
- Was the package too densely filled, tightly wrapped, or oversized for the selected system?
- Were sharp points protected without creating unnecessary bulk or stress on the package?
- Was the internal chemical indicator positioned as directed and, where applicable, in a location that provides useful information about penetration?
- Was the package loaded according to the applicable instructions?
These questions do not replace cycle validation, process monitoring, or manufacturer instructions. They provide a structured way to identify configuration drift and distinguish a packaging question from a broader process question.
What monitoring can—and cannot—show
Mechanical, chemical, and biological monitoring provide different information about sterilization. Chemical indicators can show exposure to specified process conditions, and an internal indicator may provide information about conditions within a package. An acceptable indicator response does not, by itself, demonstrate that the package will remain intact during storage or transport. CDC guidance states that chemical indicators do not by themselves guarantee sterilization and should be interpreted with mechanical and biological monitoring. [3]
The reverse is also important: a visually intact package does not prove that every aspect of sterilant access was adequate. Package inspection, process records, monitoring results, and adherence to the instructed configuration answer different questions. One observation should not be used as a substitute for the others.
Investigating a package-related event
If a package is wet, torn, open, punctured, stained, or associated with an unexpected indicator result, place the item on hold according to facility procedure and document the available facts. Inspection and investigation should address the damaged package directly rather than relying only on the indicator result. [1][2]
- device type, package type, and packaging lot or batch information when available;
- the person or team that prepared and sealed the package;
- seal or wrap appearance and the observed defect;
- sterilizer, cycle, load number, and load configuration;
- internal and external chemical-indicator results;
- mechanical records and biological-monitoring results, where applicable;
- handling, transport, storage, and event history after processing; and
- whether similar packages from the same load or packaging batch show the same problem.
The FDA educational source discusses package-integrity and stability testing in relation to stresses associated with manufacturing, sterilization, distribution, and storage. Its primary context is validation and testing of finished packaging for terminally sterilized medical devices. It is not a source for routine CSSD package-release criteria and should not be treated as a comprehensive validation standard for reusable CSSD packaging. [5]
The source of a failure should not be assumed without evidence. A defect may arise during packaging, processing, handling, transport, or storage. Investigation should therefore compare package condition with process records and the history of the item after sterilization.
Reducing avoidable variation
Quality teams can reduce variation by defining acceptance criteria for package preparation and inspection, training staff on the specific packaging systems in use, and reviewing recurring defects by package type and device family. Trend reviews should distinguish seal failures, material damage, wetness, indicator abnormalities, and storage-related damage rather than grouping every event as a generic packaging issue.
When a new package, device, sterilizer, or load configuration is introduced, evaluate the change under the facility’s established procedures. CDC recommendations call for chemical and biological testing when major changes are made to packaging, wraps, or load configuration, with processed items held during the evaluation period. [3]
Limitations
This framework does not establish cycle parameters, package shelf lives, acceptance limits, or validation protocols. ISO 11607, ISO/TS 16775, and the FDA package-integrity discussion cited here concern packaging systems for terminally sterilized medical devices and should not be read as universal requirements for every reusable CSSD packaging scenario. Requirements depend on the device, packaging system, sterilizer, applicable instructions, facility procedures, and governing requirements.
Package inspection cannot substitute for process validation, and a monitoring result cannot replace investigation of a damaged or improperly sealed package. For questions specifically involving package–sterilizer compatibility or event-related sterile-barrier inspection, consult the applicable dedicated guidance in addition to this focused framework.
Educational disclaimer
This article is for professional education and quality-improvement discussion. It is not patient-specific medical advice and does not replace device, packaging, sterilizer, regulatory, or facility instructions.
References
- Sterilizing Practices. Centers for Disease Control and Prevention. 2023-11-28.
- Guidelines in Practice: Sterilization Packaging Systems. AORN Journal via PubMed. 2020-09.
- Recommendations for Disinfection and Sterilization in Healthcare Facilities. Centers for Disease Control and Prevention. 2024-06-27.
- Recognized Consensus Standards: Medical Devices—Packaging for Terminally Sterilized Medical Devices. U.S. Food and Drug Administration. 2023-10-17.
- Medical Device Sterilization Town Hall: Sterilization Short Topics and Open Q&A. U.S. Food and Drug Administration. 2024-08-07.
Educational overview. Follow the applicable product instructions, validated procedures, local regulations and your facility’s approved policies.