Low-Temperature Sterilization: Why Load Configuration Is a Validation Variable
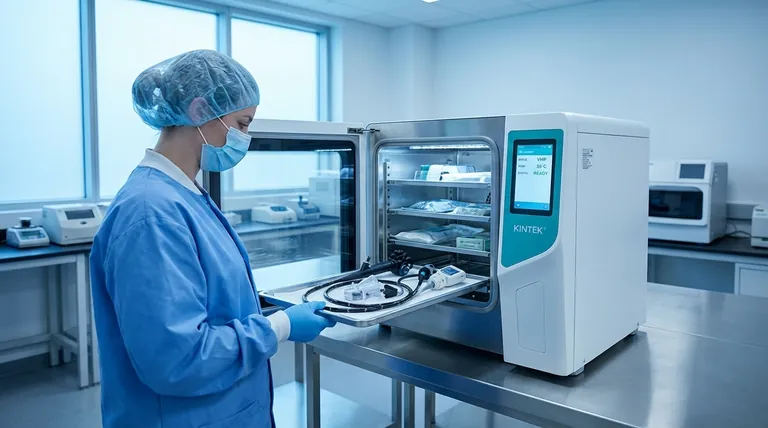
Low-temperature sterilization is often selected for devices that may be damaged by steam or dry heat. Selecting a modality, however, is only one part of sterilization assurance. Device design, packaging, materials, lumen characteristics, load arrangement, and the selected cycle must be considered together within the applicable validated process.
For central sterile services departments (CSSDs) and device-manufacturing teams, one practical lesson is especially important: load configuration is a process variable. A routine cycle result cannot automatically demonstrate that every different arrangement of devices received the same sterilization exposure or remains within the qualified process scope.
Why load configuration matters
The suitability of a sterilization process depends on the relationship between the modality and the device, materials, packaging, and areas that the sterilant must reach. The U.S. Food and Drug Administration describes several sterilization modalities used for medical devices and emphasizes that the appropriate method depends on the device and its characteristics. That general information does not, by itself, establish the performance of a particular vaporized hydrogen peroxide (VH2O2) cycle for a specific device geometry, packaging system, or challenging location.[1] (FDA)
For VH2O2 processes, the arrangement of items may change the available space, the exposure of surfaces, and the interaction between the process, the load, and materials that can affect hydrogen peroxide. A peer-reviewed review of eight qualification reports found that load configuration was specified in the reports, while load size and mass were not mentioned. Those observations describe the eight reports reviewed and should not be treated as a universal description of all VH2O2 qualification activities.[2] (PubMed)
This evidence supports a cautious conclusion: a cycle printout or indicator result is not a substitute for documented process development and qualification. Conclusions about a particular load must remain tied to the applicable sterilizer, cycle, device or device family, packaging system, loading pattern, and validation evidence.
Separate three questions that are often mixed together
1. Was the sterilizer cycle completed?
Mechanical and chemical monitoring can provide information about the cycle and the exposure conditions at the monitoring location. These data are useful, but they answer a limited question: whether the process produced the expected recorded or indicator response under the conditions observed.
2. Was the specific load within the validated scope?
This is a different question. The load may include a particular device family, packaging system, number of trays, lumen arrangement, or material combination. If the actual load differs from the qualified configuration, routine monitoring may not establish that the changed arrangement received the same validated treatment.
Process development and validation should address the load characteristics relevant to the process being used. The FDA provides general information about sterilization modalities for medical devices; the applicable device, sterilizer, organizational procedures, and supporting validation evidence determine how a specific process is developed, qualified, controlled, and released.[1] The available evidence should be used to define the boundaries of approved load families rather than assuming that a cycle name alone establishes equivalent performance for all loads.
3. Was the device and packaging system compatible with the process?
Compatibility involves more than visible damage. Depending on the process and the device, teams may need to consider packaging integrity, material effects, residues, functional performance, and whether the sterilant can reach relevant surfaces. WHO guidance on reprocessing medical devices addresses compatibility between packaging systems and sterilization methods, including the need for packaging to support sterilant access and removal while maintaining integrity through processing and handling.[3] (WHO)
That general guidance should not be used as a device-specific qualification. Packaging selection and performance still need to be evaluated against the applicable sterilizer instructions, device instructions, packaging instructions, and validation evidence.
Practical controls for CSSD and manufacturing teams
Define the load family
Describe the load in operational terms rather than relying only on a cycle name. Depending on the organization’s validated procedures, a load-family description may include:
- device type and representative challenging configuration;
- presence, length, diameter, and number of lumens or other difficult-to-access areas;
- tray, pouch, container, or other packaging system;
- quantity, mass, and arrangement of items;
- materials that may absorb, obstruct, or otherwise influence the sterilant; and
- the sterilizer, cycle, and approved loading pattern.
The level of detail should be proportionate to the process and the organization’s validated procedures. The objective is not unnecessary paperwork; it is to make changes that could affect process performance visible and reviewable.
Use approved loading diagrams or equivalent instructions
Loading instructions should show where items belong, what must remain separated, and which combinations are not permitted. If a new device, tray, packaging configuration, or accessory is introduced, treat the change as a controlled evaluation rather than an informal substitution.
CDC guidance provides general sterilization-loading principles, including placing items correctly and loosely enough to avoid impeding sterilant penetration. These principles are not modality-specific evidence for VH2O2 or another low-temperature process. They must be confirmed against the applicable sterilizer and device manufacturers’ instructions, packaging instructions, validated procedures, and supporting qualification evidence.[4] (CDC)
Make deviations visible at load release
A release review should identify whether the load matched the approved configuration. Useful prompts include:
- Was the load type authorized for this cycle?
- Were the quantity and arrangement within the defined limits?
- Were all packaging materials approved for the modality?
- Were there unusual empty spaces, overloading, or tightly nested items?
- Did the operator document any deviation and its disposition?
If the answer to one of these questions is no or unknown, the appropriate response depends on the organization’s validated procedure and risk assessment. It may require holding the load, consulting the device or sterilizer manufacturer, opening a deviation, or performing a documented technical assessment. A positive routine indicator should not be used to dismiss an unassessed configuration change.
How monitoring fits into the control strategy
Biological and chemical indicators can be important components of routine control when used according to the applicable process and instructions. However, indicator performance, placement, and interpretation are modality- and manufacturer-specific. Monitoring devices challenge or measure only the locations and conditions for which they are designed; they do not automatically represent every device geometry, packaging system, or load arrangement.
Literature on parametric release, including discussion of ethylene oxide and VH2O2, emphasizes that process data are most informative when the load has already been addressed through validation. Load variability that was not included in performance qualification may limit what biological indicators, chemical indicators, or process measurements can demonstrate.[5] (PubMed) This is a literature-based warning about the relationship between data interpretation and validated load scope, not a universal statement that every process must use the same routine-release approach.
Limitations and escalation points
There is no universal loading pattern that can be transferred between sterilizers, cycles, device families, or packaging systems. A configuration that performs acceptably in one validated process may not be suitable in another. Manufacturer instructions, applicable standards, local procedures, and the organization’s validation evidence remain controlling.
Escalate when a proposed change involves a new device design, new packaging, increased quantity or mass, altered lumen characteristics, a different sterilizer, a different cycle, or a recurring monitoring anomaly. The need for additional qualification or testing cannot be determined from a generic article alone.
Key takeaway
For low-temperature sterilization, “the cycle ran” and “the load was processed within the validated process” are not interchangeable statements. Treat load configuration as a controlled variable, define the boundaries of each load family, document deviations, and use routine monitoring as one part of—not a replacement for—validated process control.
Educational disclaimer: This article provides general educational information for sterilization and sterile-processing professionals. It does not replace device-specific instructions for use, sterilizer instructions, applicable standards, validation records, regulatory requirements, or qualified technical assessment.
References
- Sterilization for Medical Devices. U.S. Food and Drug Administration. 2024-11-26.
- Qualification of low temperature vaporized hydrogen peroxide sterilization: Towards greater harmonization of practices?. PubMed / peer-reviewed literature. 2022.
- Decontamination and reprocessing of medical devices for health-care facilities. World Health Organization. 2016.
- Sterilizing Practices. Centers for Disease Control and Prevention. 2024.
- Principles of Parametric Release: Emphasis on Data Collection and Interpretation. PubMed / peer-reviewed literature. 2023.
Educational overview. Follow the applicable product instructions, validated procedures, local regulations and your facility’s approved policies.